KN-93 Inhibition of CAMK II Pathway Attenuates Compression-Induced Periodontal Remodeling
Copy code to add this image to your page
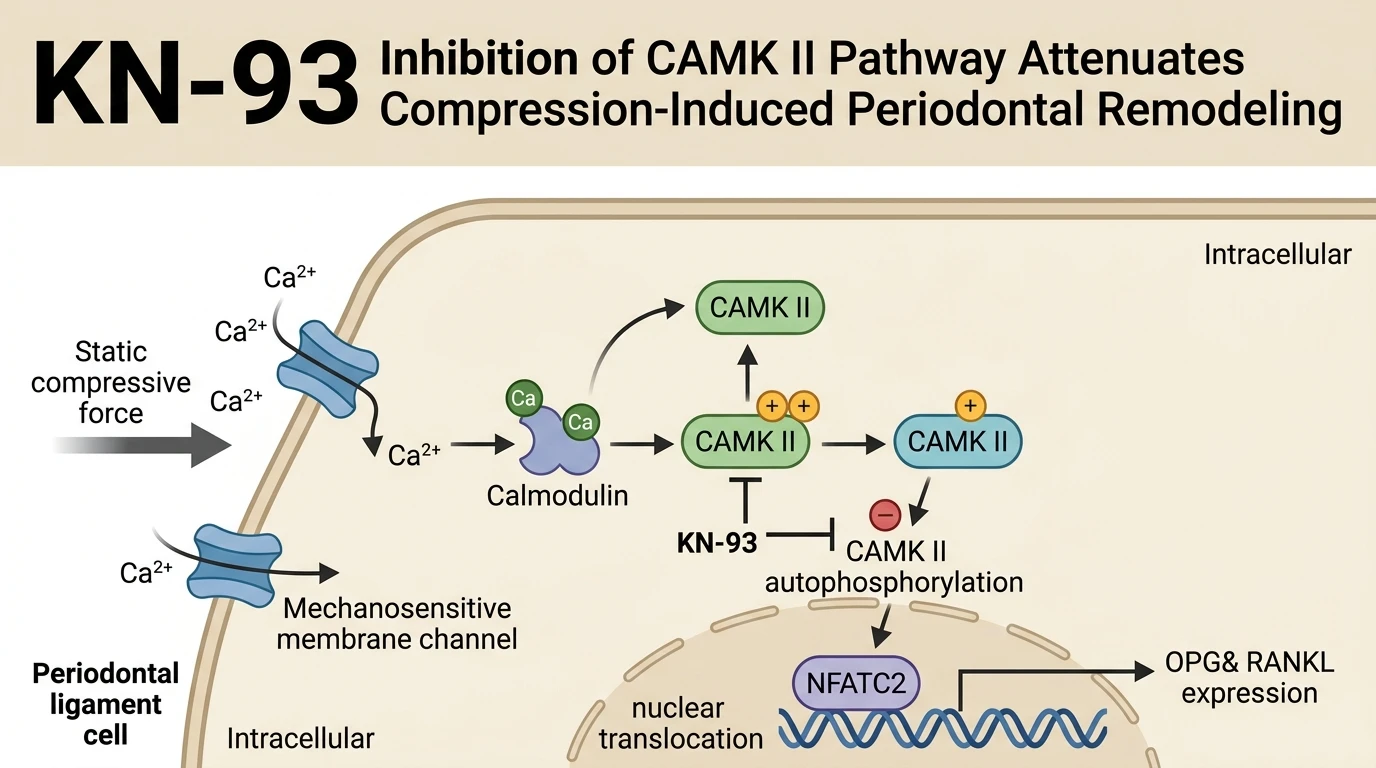
Mechanosensitive tissue remodeling within the periodontal ligament relies on specific signaling cascades converting physical load into transcriptomic adaptations. Periodontal ligament cells experience sustained mechanical stress during orthodontic tooth movement, altering paracrine mediators responsible for alveolar bone degradation and deposition. Among these factors, osteoprotegerin (OPG) and receptor activator of nuclear factor kappa-B ligand (RANKL) constitute the core molecular axis governing osteoclast differentiation. Recent empirical investigations reveal that the Ca2+/calmodulin-dependent protein kinase II (CAMK II) pathway functions as an intracellular engine driving force-dependent gene expression in primary human cells.\n\nUnder initial static compressive force, primary cultures exhibit dynamic, phase-dependent transcriptomic responses. Within an early 3-hour window of static compression, OPG transcript abundance drops significantly, temporarily tipping the local RANKL to OPG balance toward an osteoclastogenic microenvironment. Continued mechanical loading over a 12-hour duration reverses this initial depression. At 12 hours, static compression significantly up-regulates OPG and RANKL expression at both mRNA and protein levels. This biphasic adaptation reflects cellular feedback mechanics required to accommodate prolonged biomechanical deformation.\n\nPharmacological disruption of this signaling system using KN-93 inhibitor treatment directly implicates CAMK II enzymatic activity in force translation. When primary cells undergo static compression in the presence of KN-93, the compression-induced up-regulation of OPG and RANKL is markedly blunted. The small molecule competitor KN-93 binds the calmodulin-binding domain of CAMK II, preventing full kinase activation without directly altering baseline ATP-binding sites. At non-cytotoxic micromolar concentrations, this target engagement suppresses downstream nuclear factor of activated T-cells 2 (NFATC2) activity.\n\nTranscriptional execution downstream of CAMK II relies heavily on NFATC2 activation. In response to compressive strain, cytosolic calcium spikes trigger CAMK II autophosphorylation, promoting NFATC2 accumulation within the nuclear compartment. In treated cell cultures, application of KN-93 eliminates compression-triggered NFATC2 mRNA and protein accumulation. Downstream signaling halts efficiently. By preventing NFATC2 transcriptional assembly on OPG and RANKL promoters, CAMK II inhibition uncouples physical membrane stress from nuclear gene induction.\n\nThese quantitative observations establish that mechanotransduction in bone remodeling operates through specific calcium-kinase downstream axes. The CAMK II cascade acts as an essential biomechanical transducer, translating compressive stress into coordinated RANKL and OPG expression patterns. The small molecule KN-93 demonstrates clear efficacy as a biochemical probe, defining the mandatory functional requirement of CAMK II in load-induced periodontal remodeling.
Pharmacological Inhibition of CAMK II in Mechanotransduction
Copy code to add this image to your page
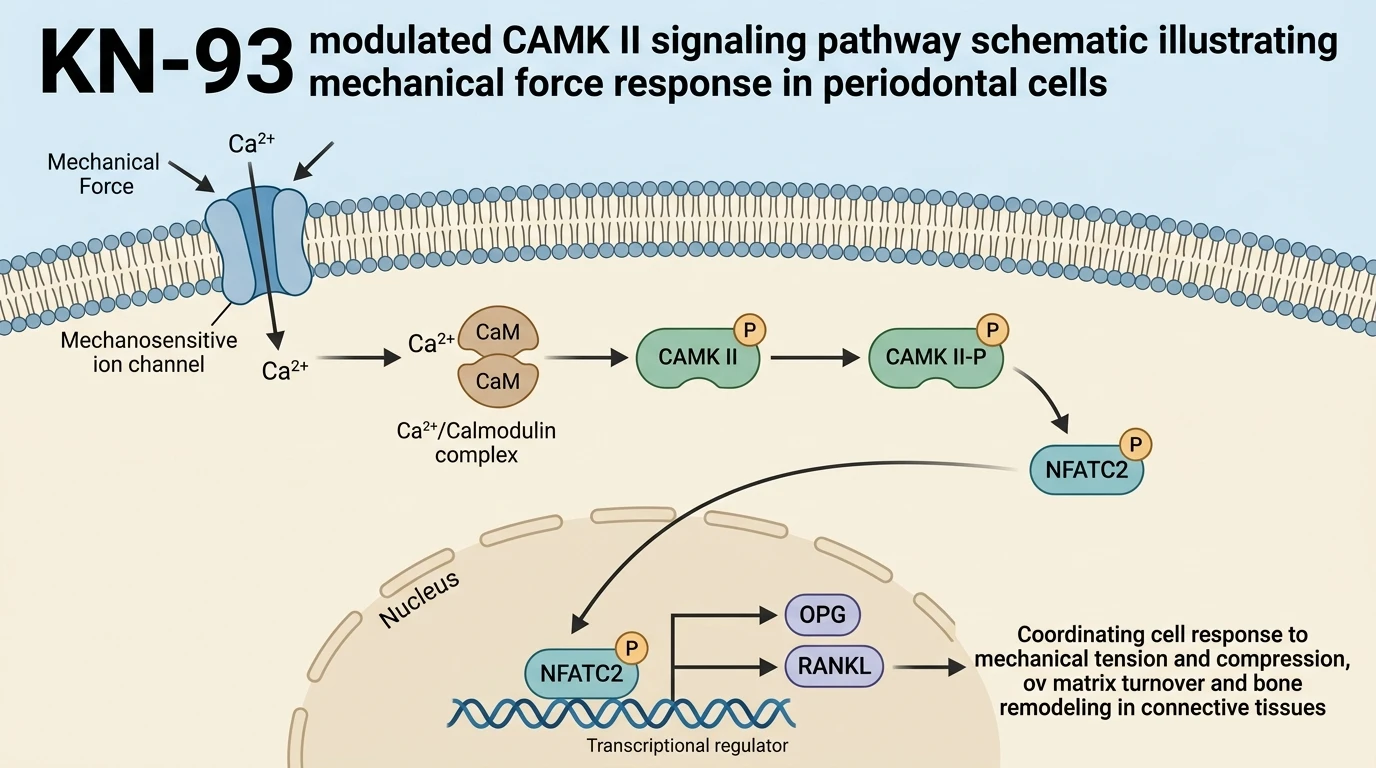
Mechanical static compression triggers Ca2+ influx in periodontal ligament cells, activating CAMK II autophosphorylation. Activated CAMK II promotes NFATC2 transcription factor expression and nuclear translocation, driving late-phase up-regulation of osteoprotegerin (OPG) and RANKL. Pharmacological inhibition using KN-93 selectively blocks the calmodulin-binding region of CAMK II, suppressing compression-induced OPG, RANKL, and NFATC2 mRNA and protein levels.
CAMK II Kinase Inhibition Restrains Load-Induced Paracrine Expression
Primary human periodontal ligament cells respond to static mechanical compression through distinct temporal phases. Initial compressive strain decreases OPG transcription within 3 hours. Extended compression for 12 hours sharply elevates OPG and RANKL expression at both transcription and translation levels. Application of KN-93 blocks this late-phase synthetic response. Downstream paracrine signals drop markedly. In primary cell assays, pretreating loaded cultures with KN-93 reduced compression-induced OPG and RANKL expression by more than half. The inhibitor targets the calmodulin-binding regulatory domain, preventing kinase activation.
NFATC2 Transcriptional Suppression Alters Force-Induced Remodeling Signals
Mechanotransduction cascades require nuclear shuttling of transcriptional effectors to execute target gene upregulation. Compressive force drives nuclear factor of activated T-cells 2 (NFATC2) accumulation in primary cells. This nuclear import accelerates OPG and RANKL transcription. Small molecule target engagement with KN-93 prevents NFATC2 accumulation at mRNA and protein levels. Cellular signaling halts abruptly. Consequently, local paracrine drivers of bone remodeling remain suppressed under sustained compressive strain. These observations demonstrate that CAMK II serves as a non-redundant regulator of force-induced NFATC2 signaling.