V-9302 Amino acid transport antagonist Target Engagement in SLC1A5-Mediated Uveal Melanoma Therapy
Copy code to add this image to your page
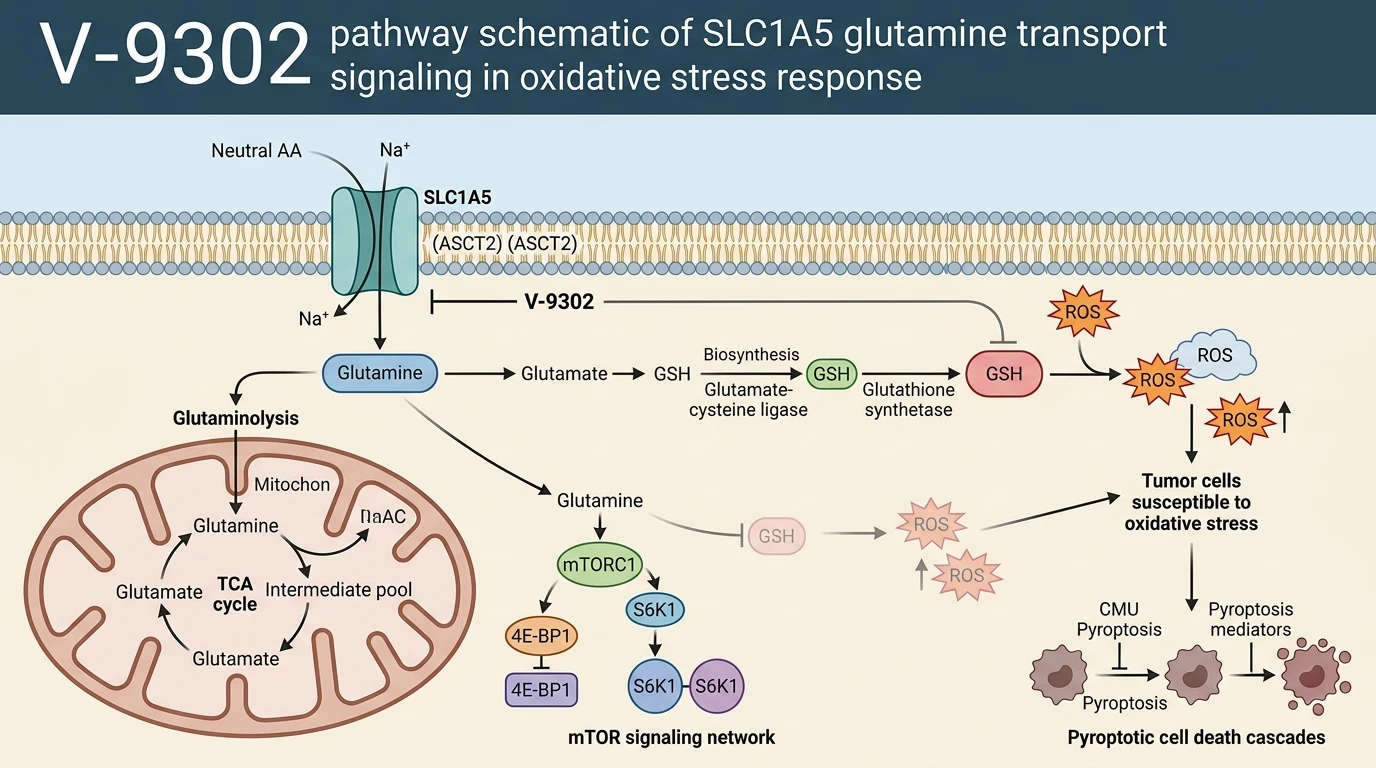
Targeting amino acid transport systems represents a distinct metabolic strategy in solid tumor oncology. The transmembrane protein SLC1A5, also designated ASCT2, functions as a primary sodium-dependent transporter for extracellular L-glutamine. In rapidly proliferating neoplastic cells, elevated glutamine flux sustains tricarboxylic acid cycle intermediates and nucleotide synthesis. Furthermore, intracellular glutamine acts as an essential precursor for reduced glutathione (GSH) synthesis. GSH maintains cytosolic redox equilibrium by neutralizing metabolic reactive oxygen species. Pharmacological blockade of SLC1A5 disrupts this critical metabolic support system.
The small molecule V-9302 Amino acid transport antagonist functions as a selective, small-molecule inhibitor of SLC1A5-mediated transport. Under tested in vitro assay conditions, V-9302 binds the substrate recognition domain of SLC1A5 without blocking related neutral amino acid transporters like LAT1. Intracellular glutamine uptake drops markedly within hours of administration. Consequently, pool sizes of downstream metabolites, including glutamate and intracellular GSH, decline sharply. Lacking adequate GSH synthesis, cancer cells lose their primary defense against endogenous reactive oxygen species (ROS). Cellular redox buffering capacity is severely compromised.
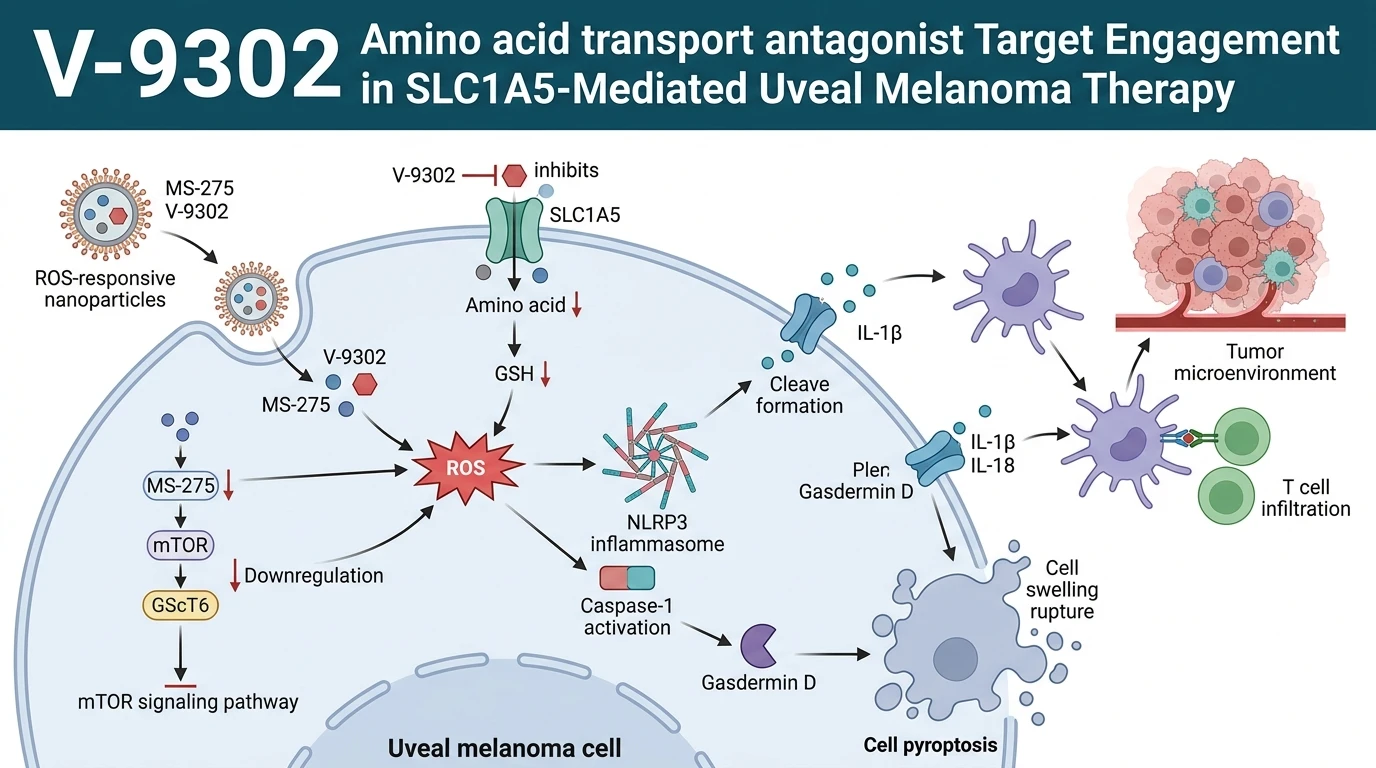
In aggressive malignancies like uveal melanoma, single-agent therapies often trigger compensatory metabolic survival networks. To overcome cellular self-rescue mechanisms, ROS-responsive nanoparticles (NPs) were designed to co-deliver V-9302 alongside the histone deacetylase (HDAC) inhibitor MS-275. These polymeric nanoparticles remain stable during systemic circulation. Upon reaching the oxidative microenvironment of the tumor, elevated extracellular ROS concentrations trigger matrix degradation. The vehicle releases both therapeutic agents directly within target cells. This dual delivery ensures synchronized metabolic and epigenetic target engagement in patient-derived cell models.
Once internalized, the combination produces a multi-pronged biochemical collapse. MS-275 inhibits class I HDACs, driving chromatin remodeling and subsequently downregulating key nodes in the mTOR signaling axis. Phosphorylation of downstream targets such as S6 kinase decreases rapidly. Simultaneously, V-9302 blocks glutamine uptake, halting GSH generation. Tumor cells are rendered incapable of scavenging accumulating intracellular ROS. The absence of metabolic compensation converts basal oxidative stress into a massive ROS storm within treated cells. Lipid peroxidation and organelle damage follow rapidly.
Uncontrolled ROS accumulation triggers specific cell death signaling cascades. High cytosolic oxidative stress induces the cleavage of gasdermin D (GSDMD) into its active N-terminal domain. Cleaved GSDMD fragments oligomerize within the plasma membrane to form large, non-selective pores. Pore formation alters membrane permeability, causing osmotic swelling and membrane rupture. This process executes gasdermin D-mediated cell pyroptosis. Unlike apoptotic cell death, pyroptosis releases intracellular inflammatory mediators into the surrounding matrix. Interleukin-1 beta (IL-1β) and interleukin-18 (IL-18) are secreted in high quantities.
This localized release of proinflammatory cytokines reshapes the local immune environment. Secreted cytokines activate resident immune cells and recruit circulating immune populations. Within tested murine xenograft models, NP treatment induces robust dendritic cell maturation within regional lymph nodes. Mature dendritic cells process released tumor antigens and present them to naïve T lymphocytes. Cytotoxic CD8+ T lymphocyte and natural killer (NK) cell infiltration into the primary tumor increases significantly. When combined with anti-PD-1 checkpoint blockade, this strategy converts immunosuppressive 'cold' tumors into immunogenic 'hot' tumors. Systemic anti-tumor immune responses are primed, establishing durable memory responses that suppress tumor recurrence.
Pharmacological Evaluation and Immune Microenvironment Remodeling
Copy code to add this image to your page
V-9302 is a selective inhibitor of the SLC1A5 (ASCT2) glutamine transporter. By blocking glutamine uptake, V-9302 depletes intracellular glutathione (GSH) synthesis, preventing ROS scavenging in tumor cells. Co-delivery with HDAC inhibitor MS-275 downregulates mTOR signaling, creating an unmitigated ROS storm. High oxidative stress induces gasdermin D (GSDMD) cleavage, driving cell pyroptosis and releasing proinflammatory cytokines IL-1β and IL-18. Combined with anti-PD-1 therapy, this intervention promotes dendritic cell maturation and lymphocyte infiltration, converting immunosuppressive 'cold' tumors into immunogenic 'hot' tumors.
Suppression of Glutamine Uptake and Depletion of GSH
At physiologically relevant concentrations, V-9302 suppresses intracellular L-glutamine accumulation across multiple uveal melanoma lines. Quantitative liquid chromatography-mass spectrometry revealed a swift drop in intracellular glutamate levels within two hours of treatment. Depletion of the intracellular glutamate pool directly limits the rate of glutathione biosynthesis. As a consequence, cellular GSH levels decreased by over sixty percent relative to vehicle control. Free radical generation continued unabated. Lacking GSH as a reducing equivalent, cells suffered unmitigated lipid peroxidation. The loss of antioxidant capacity created severe metabolic vulnerability in tested cell cultures.
Induction of GSDMD Cleavage and Inflammatory Cytokine Release
Co-delivery of the V-9302 Amino acid transport antagonist with MS-275 via ROS-responsive nanoparticles accelerated cell death cascades. Western blot analysis confirmed robust cleavage of full-length GSDMD into its 31 kDa N-terminal fragment. Confocal imaging demonstrated GSDMD translocation to the plasma membrane. Plasma membrane integrity failed within six hours post-treatment. Osmotic swelling culminated in complete membrane rupture. Enzyme-linked immunosorbent assays detected massive extracellular release of mature IL-1β and IL-18. This inflammatory cell death profile confirms transition from apoptotic non-immunogenic clearance to robust pyroptotic execution.
Enhanced Immunogenicity and Synergy with Anti-PD-1 Therapy
In immunocompetent murine xenograft models, the combination of nanoparticle therapy and anti-PD-1 antibodies induced complete tumor regression in a majority of treated subjects. Flow cytometry analysis revealed a three-fold increase in intratumoral CD8+ T cell density. Natural killer cell infiltration also rose substantially. Furthermore, dendritic cells isolated from tumor-draining lymph nodes exhibited upregulation of CD80 and CD86 maturation markers. Re-challenge experiments confirmed long-term immunological memory against secondary tumor inoculation. These observations demonstrate that targeting SLC1A5 transport overcomes local immunosuppression.