MIK665 (S64315) Inhibits MCL-1 to Restore Apoptosis in Venetoclax-Resistant Myeloid Malignancies
Copy code to add this image to your page
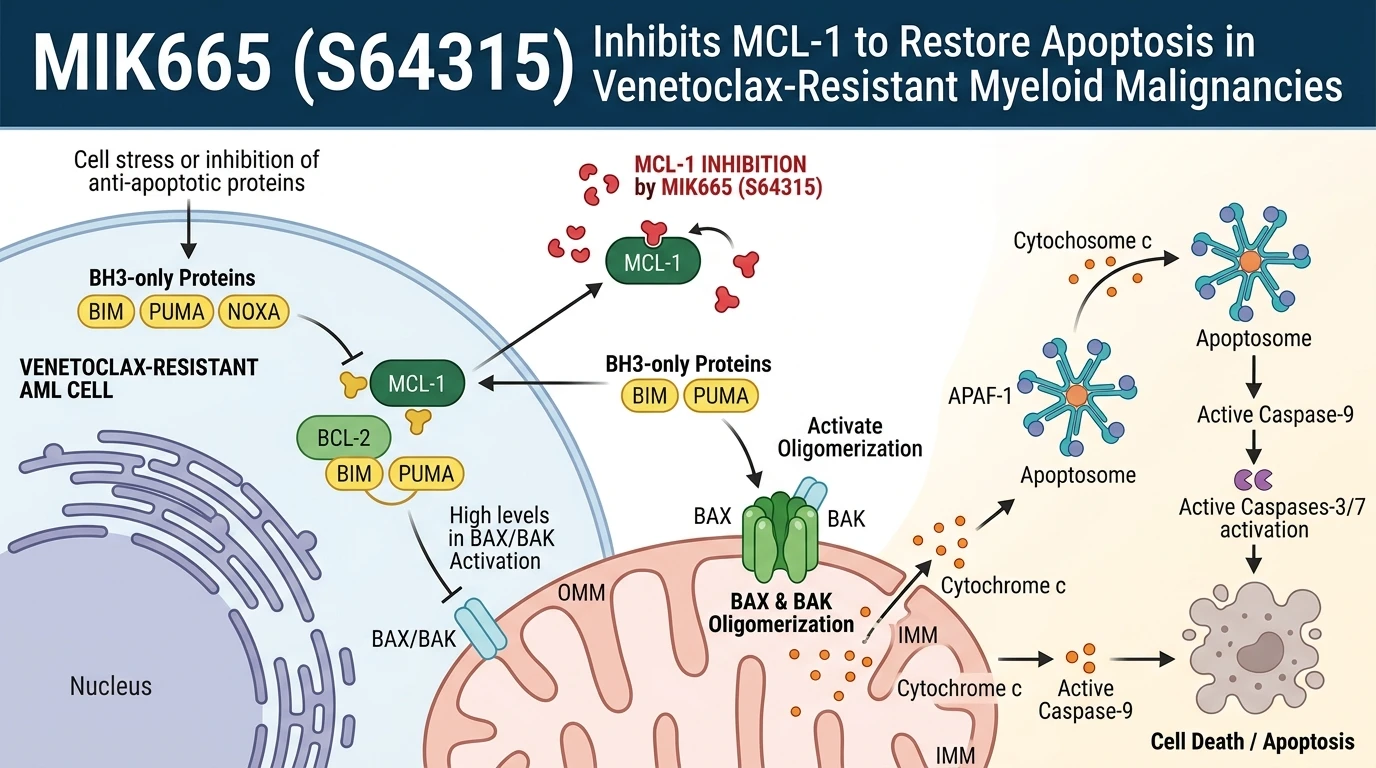
Evasion of programmed cell death represents a hallmark of hematologic malignancies, particularly myelodysplastic syndromes (MDS) and acute myeloid leukemia (AML). Central to this survival phenotype is the regulation of the intrinsic mitochondrial apoptosis pathway by the B-cell lymphoma 2 (BCL-2) family of proteins. This family is divided into pro-survival factors—such as BCL-2, BCL-XL, and myeloid cell leukemia 1 (MCL-1)—and pro-apoptotic effectors, including BAX, BAK, and BH3-only sensitizers like BIM, NOXA, and PUMA. Under physiological conditions, MCL-1 neutralizes pro-apoptotic activity by binding tightly to the alpha-helical BH3 domains of BAX, BAK, and activator BH3-only proteins, preventing executioner protein oligomerization. In dysplastic myeloid blasts, MCL-1 is frequently overexpressed, suppressing apoptotic cascades and providing a crucial survival buffer against cytotoxic stress.
The clinical validation of BCL-2 inhibition via small molecules such as venetoclax marked a major advance in hematologic oncology. However, primary and acquired resistance to selective BCL-2 blockade remains a widespread challenge in AML management. A primary mechanism driving this resistance involves the compensatory upregulation or reliance on MCL-1. When BCL-2 is pharmacologically inhibited, pro-apoptotic proteins such as BIM are displaced from BCL-2 and subsequently sequestered by MCL-1. This compensatory shift creates an effective bypass pathway, enabling malignant cells to evade mitochondrial outer membrane permeabilization (MOMP) despite complete BCL-2 occupancy. Consequently, targeted inhibition of MCL-1 has emerged as an essential therapeutic strategy to re-establish apoptotic sensitivity in resistant cell populations.
MIK665 (S64315) was engineered as a potent, highly selective, small-molecule BH3 mimetic designed to address this therapeutic gap. Structurally, S64315 docks into the hydrophobic BH3-binding groove of MCL-1 with nanomolar affinity, mimicking the endogenous interactions of native BH3-only proteins. X-ray crystallography and biophysical binding assays demonstrate that S64315 occupies the p2 to p4 hydrophobic pockets within the BH3-binding cleft of MCL-1, forming key electrostatic interactions and salt bridges that stabilize the complex. Crucially, the compound exhibits high selectivity for MCL-1 over closely related paralogs BCL-2 and BCL-XL, minimizing off-target inhibition that could induce systemic toxicities such as thrombocytopenia. Upon engagement with MCL-1, S64315 sterically displaces bound BIM, NOXA, and effector proteins BAX and BAK.
The competitive displacement of these pro-apoptotic molecules triggers the execution phase of intrinsic apoptosis. Unbound BAX and BAK undergo conformational changes, exposing their N-terminal domains and driving homooligomerization within the outer mitochondrial membrane. This process generates supramolecular pores that disrupt membrane integrity, leading to MOMP. Following MOMP, key apoptogenic factors, including cytochrome c and second mitochondria-derived activator of caspases (SMAC/DIABLO), diffuse from the intermembrane space into the cytosol. Cytosolic cytochrome c complexes with apoptotic protease activating factor 1 (APAF-1) and pro-caspase-9 in an ATP-dependent manner to assemble the apoptosome. Activator caspase-9 subsequently cleaves and activates executioner caspases-3 and -7, initiating widespread proteolytic cleavage of structural and functional cellular substrates, ultimately terminating in apoptotic cell death.
In preclinical models of MDS and AML, treatment with MIK665 (S64315) demonstrates pronounced single-agent cytotoxicity across primary blast samples harboring diverse cytogenetic profiles. Pharmacodynamic analyses in patient-derived blast models confirm that S64315 induces rapid caspase-3 cleavage and PARP proteolysis within four to six hours post-administration. The degree of cell death strongly correlates with baseline MCL-1 dependency rather than total protein expression levels alone, highlighting the importance of functional priming in predicting drug sensitivity. Furthermore, rational drug combination studies demonstrate that combining S64315 with venetoclax concurrently blocks both BCL-2 and MCL-1 survival nodes, effectively eliminating the rescue mechanism that drives resistance. In translational murine xenograft studies, intermittent dosing schedules of S64315 achieved robust tumor regression while allowing transient recovery of physiological cell populations dependent on MCL-1 survival signaling. These preclinical findings provided the rationale for evaluating MIK665 (S64315) in Phase I clinical trials for patients with relapsed or refractory MDS and AML, where target engagement, safety profiles, and preliminary pharmacodynamic biomarkers continue to be systematically defined.
Pharmacological Mechanisms and Efficacy in Hematologic Malignancies
Copy code to add this image to your page
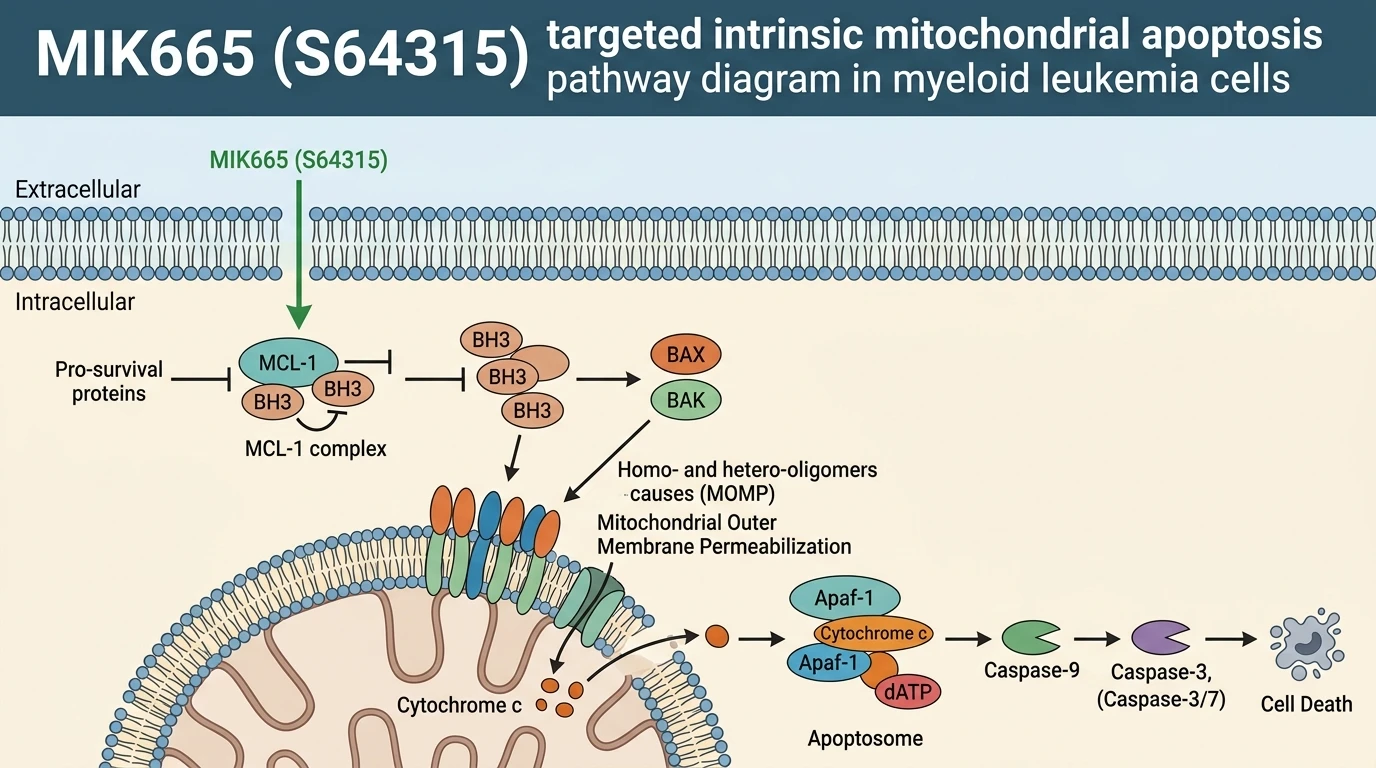
MIK665 (S64315) functions as a potent BH3 mimetic that selectively binds the anti-apoptotic protein MCL-1. By competitive binding to the hydrophobic BH3 groove, it prevents MCL-1 from sequestering pro-apoptotic BH3-only proteins like BIM. The resulting liberation of pro-apoptotic effectors permits BAX and BAK oligomerization on the mitochondrial outer membrane, inducing membrane permeabilization, cytochrome c release, and subsequent caspase cascade activation. This targeted inhibition effectively circumvents MCL-1-mediated venetoclax resistance in dysplastic myeloid cells.
Displacement of Pro-Apoptotic Factors Triggers MOMP Assembly
In human acute myeloid leukemia blast isolates, selective engagement of the MCL-1 hydrophobic binding groove by S64315 disrupts high-affinity interactions with BIM and NOXA. Free pro-apoptotic BH3-only proteins subsequently activate executioner proteins BAX and BAK on the outer mitochondrial membrane; this event induces oligomerization and subsequent mitochondrial outer membrane permeabilization (MOMP). In preclinical cell-free assays, mitochondrial membrane potential loss occurred within two hours of treatment. Cytosolic cytochrome c accumulation followed rapidly. These observations establish that S64315 initiates apoptosis via direct mitochondrial destabilization rather than off-target cytotoxicity.
Combination Efficacy Overcomes Venetoclax Resistance in Blast Models
Upregulation of anti-apoptotic MCL-1 represents a primary bypass pathway responsible for acquired resistance to the selective BCL-2 inhibitor venetoclax. Co-administration of S64315 effectively neutralizes this survival adaptation in patient-derived dysplastic myeloid cells. Combined targeting of BCL-2 and MCL-1 restored BAX/BAK activation under in vitro culture conditions, demonstrating synergistic lethality against resistant blast populations. In murine xenograft models of refractory acute myeloid leukemia, dual blockade yielded pronounced tumor regression without inducing overt systemic toxicity. Preliminary Phase I clinical evaluation remains ongoing to establish tolerable dosage regimens and pharmacodynamic markers in patients with high-risk myelodysplastic syndromes.